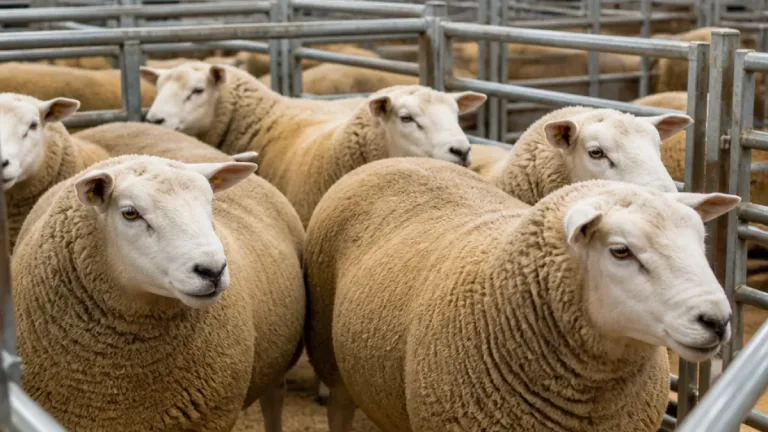
Researchers have identified a missense variant in the INHA gene that is strongly associated with litter size in a flock of 100 Kreishi ewes, according to a study published in Molecular Biology Reports. The team, led by Siraj A. Hashem, Mervan Bayraktar and Tahreer M. Al-Thuwaini, found a cytosine-to-thymine substitution at nucleotide position 425 (c.425C>T) that changes amino acid 142 from serine to phenylalanine (p.Ser142Phe).
Study population and methods
The study examined reproductive records for ewes aged three to four years and split the flock into two groups based on lambing outcome: 46 ewes that had delivered single lambs and 54 ewes that had delivered twins. Researchers genotyped the INHA site by polymerase chain reaction followed by bidirectional Sanger sequencing and observed three genotypes in the flock, designated CC, CT and TT.
Statistical findings
Genotype distribution differed markedly between single-bearing and twin-bearing ewes. A chi-square test produced a statistic of 30.87 with a P value < 0.0001, indicating a very low probability that the observed association arose by chance. Binary logistic regression quantified the associations: compared with ewes homozygous for the TT genotype, animals with the CT genotype had an odds ratio of 6.77 (95% confidence interval 1.59–28.72) for giving birth to twins. The CC genotype showed a larger estimated effect, with an odds ratio of 16.51 (95% confidence interval 5.56–49.08).
The authors emphasise that these estimates, especially for the CT group, have wide confidence intervals, reflecting statistical uncertainty arising from smaller genotype subgroup sizes. They note that the result demonstrates a statistical association in a single flock rather than proving a causal biological mechanism, and they call for replication in larger, independent populations before the variant could be used as a molecular marker for breeding selection.
- Study size: 100 Kreishi ewes (aged three to four years)
- Reproductive groups: 46 single-bearing, 54 twin-bearing
- Variant: c.425C>T in INHA, resulting in p.Ser142Phe
- Key stats: chi-square 30.87, P < 0.0001; OR CT v TT 6.77 (95% CI 1.59–28.72); OR CC v TT 16.51 (95% CI 5.56–49.08)
| measure | value |
|---|---|
| total ewes | 100 |
| single-bearing | 46 |
| twin-bearing | 54 |
| chi-square | 30.87 |
| P value | < 0.0001 |
Implications and cautions
The INHA gene is a biological candidate for reproductive traits, and the p.Ser142Phe substitution reported here is plausibly functional, but the authors stop short of asserting causation. They recommend validation work in larger and genetically distinct flocks before the variant is considered for use as a selection marker in breeding programmes. The paper also highlights the limitations of single-population association studies and the need to quantify any potential trade-offs or pleiotropic effects before applying findings in commercial selection schemes.
Until such validation is completed, breeders and advisers should treat the finding as an important lead for further research rather than as a ready-to-use genetic test. The study's authors and the journal are cited in the published report for full methodological details and interpretation.